Navigating the Approval Gap: How Forward-Thinking Biotech Companies Are Outmaneuvering a Drug Development System Built for a Different Era
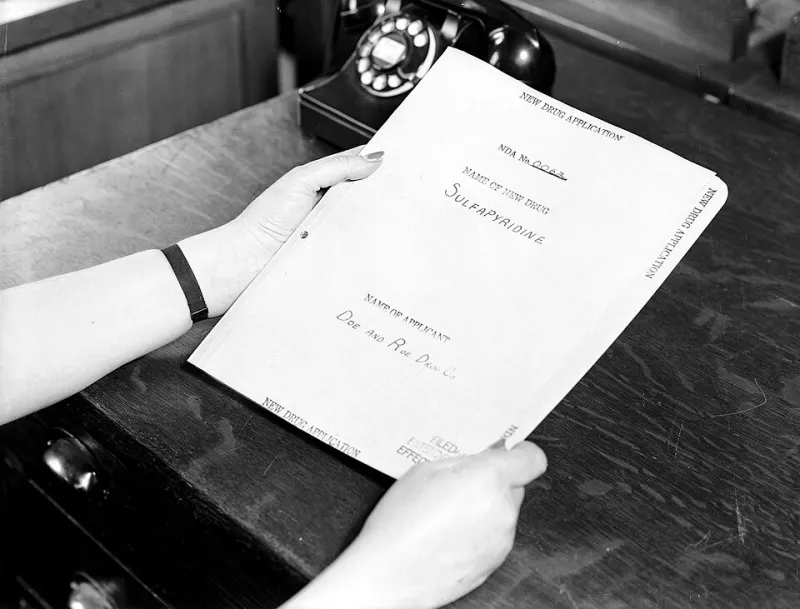
Photo: The U.S. Food and Drug Administration, Public domain, via Wikimedia Commons
The average time from initial compound identification to FDA approval for a new drug in the United States sits somewhere between ten and fifteen years. Depending on the therapeutic area, the figure can be considerably longer. For patients waiting on treatments for serious or rare conditions, that timeline is not an abstraction — it is a lived reality with measurable human cost. For the biotech companies bearing the financial and operational burden of that journey, it represents a structural constraint that shapes every strategic decision from early research through commercialization.
The tension at the center of this dynamic is genuine and not easily resolved. Regulatory rigor exists for sound reasons. The history of pharmaceutical development includes episodes — some catastrophic — that illustrate exactly why independent, evidence-based oversight is essential. The FDA's mandate to ensure that drugs are both safe and effective before reaching the public is not a bureaucratic preference; it is a foundational principle of modern medicine.
And yet the mechanisms through which that mandate is executed were designed in a different era of science. They were built around assumptions about how diseases work, how evidence is generated, and how clinical trials are structured that no longer fully reflect the state of the field. The friction this creates is real, and the biotech companies navigating it most successfully are doing so not by circumventing oversight, but by engaging with the regulatory system in more sophisticated ways.
Where the System Creates Unintended Friction
The traditional drug approval pathway — preclinical work, Phase I through Phase III clinical trials, New Drug Application, review — was designed around small-molecule compounds targeting broadly defined patient populations. It was built to evaluate interventions that are relatively uniform in their mechanism and whose effects can be measured across large, randomized cohorts over extended periods.
Precision medicine has complicated this model considerably. When a therapeutic is designed to work in a patient population defined by a specific genetic mutation or biomarker profile, the relevant patient group may number in the thousands nationally. Recruiting sufficient trial participants under traditional statistical frameworks becomes prohibitively difficult. The evidentiary standards that made sense for a blockbuster cardiovascular drug do not translate cleanly to an orphan disease treatment or a cell therapy designed for a molecularly defined subpopulation.
The challenge is compounded by the pace at which underlying science is advancing. In oncology, for example, the understanding of tumor biology, resistance mechanisms, and immunological response is evolving rapidly enough that a clinical trial designed at the start of a development program may be measuring the wrong endpoints by the time it completes enrollment. The regulatory framework's built-in timelines can inadvertently lock companies into scientific assumptions that have since been superseded.
The Pathways That Are Changing the Equation
The FDA has, to its credit, developed a range of expedited and adaptive mechanisms that sophisticated biotech companies are now using as primary strategic tools rather than secondary options.
Breakthrough Therapy Designation, established in 2012, allows the agency to provide intensive guidance to developers of drugs that show preliminary clinical evidence of substantial improvement over existing treatments for serious conditions. Companies that secure this designation report meaningfully more productive interactions with FDA reviewers — interactions that can surface and resolve potential approval obstacles years earlier in the development process than traditional review would allow.
Adaptive trial designs represent another significant lever. Rather than committing to a fixed protocol from enrollment to completion, adaptive trials allow predefined modifications — adjustments to dosing, patient stratification, or endpoint weighting — based on interim data. When executed with appropriate statistical rigor and pre-specified decision rules, these designs can reduce both the time and the patient exposure required to generate a statistically valid result. The FDA's Complex Innovative Trial Design program has been an important vehicle for advancing this approach.
Real-world evidence is perhaps the most consequential emerging element of the regulatory toolkit. Data generated outside the controlled clinical trial environment — from electronic health records, insurance claims databases, patient registries, and wearable devices — is increasingly being recognized as a legitimate complement to traditional trial data, particularly for post-approval commitments and label expansions. Companies that invest in the infrastructure to collect and analyze real-world data are building a regulatory asset that extends well beyond initial approval.
The Strategic Dimension of Regulatory Engagement
The most important shift in how leading biotech companies approach the regulatory process is attitudinal: they are treating FDA engagement as a core competency rather than a downstream obligation.
This means investing in regulatory affairs expertise at the earliest stages of program design, before clinical development begins. It means structuring development programs around the evidentiary questions the agency is likely to ask, rather than the questions that are most convenient to answer. It means maintaining an ongoing dialogue with the agency through pre-IND meetings, Type B and Type C meeting requests, and participation in pilot programs designed to test new approval frameworks.
Companies that approach this engagement proactively tend to encounter fewer late-stage surprises — the complete response letters and unexpected information requests that can add years to a program's timeline. They also tend to build institutional knowledge that compounds over successive programs, creating a durable organizational capability that smaller, less experienced competitors cannot easily replicate.
Balancing Speed With Rigor
Any serious analysis of regulatory reform in drug development must grapple with the legitimate concerns on the other side of the ledger. Accelerated approvals have, in some cases, been granted for therapies that subsequent confirmatory trials have shown to be less effective than surrogate endpoints suggested. The pressure to move quickly is real, but so is the risk of moving prematurely.
The answer is not a binary choice between speed and safety. It is the development of more sophisticated evidentiary frameworks — ones that can distinguish between genuine scientific uncertainty and administrative delay, that can incorporate diverse data sources without sacrificing analytical rigor, and that can evolve in step with the science they are designed to evaluate.
The biotech companies contributing most constructively to this evolution are those that engage with the regulatory system not as an obstacle to be minimized, but as a partner in the shared project of bringing effective, safe treatments to patients as efficiently as science and evidence allow. That orientation — collaborative, rigorous, and forward-looking — is ultimately what the approval gap demands.